An LC-MS/MS peak area that climbs across an injection sequence is a pattern, not a diagnosis. The rise can come from the analyte, sample preparation, the matrix, the flow path, the ion source, detector response, or the relationship between one injection and the next. The fast route to the cause is to characterize the trend, compare analyte and internal-standard behavior, test whether it appears in neat standards or only in matrix, and then run targeted sequence experiments. Do not correct the method until an experiment distinguishes the mechanism.
How do you recognize the sequence pattern before assigning a cause?
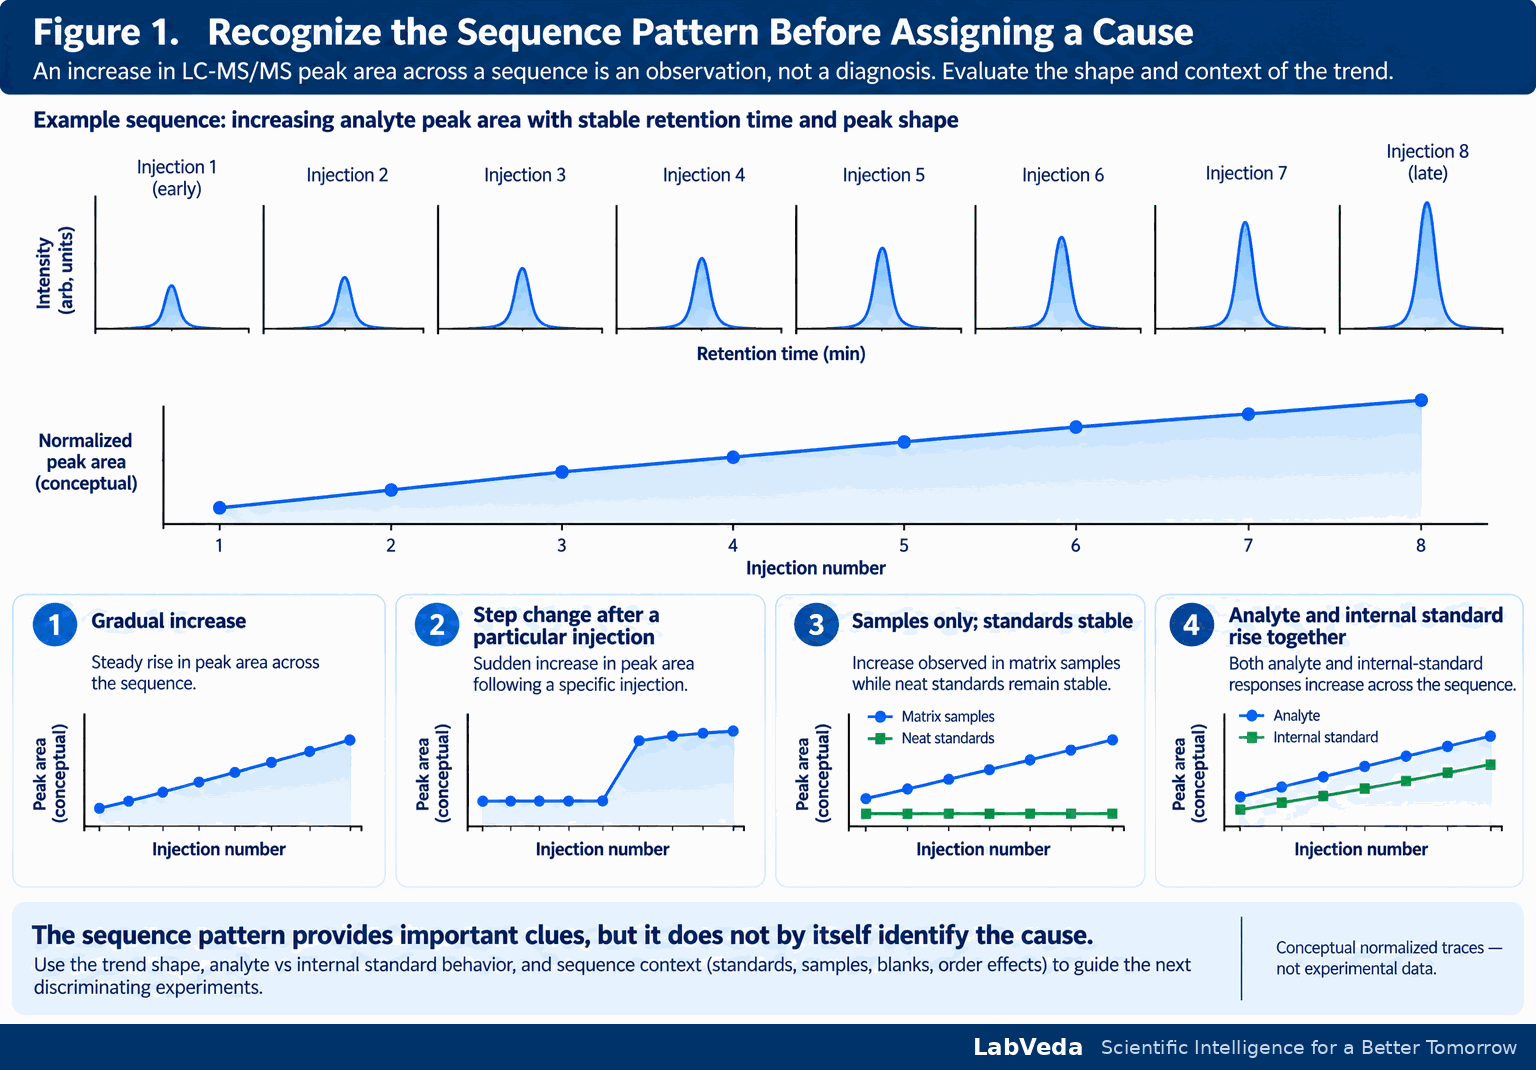
The trend may be gradual, may approach a plateau, may appear only after a particular high-response injection, or may occur only in matrix-containing samples — and these shapes are diagnostically different. Review retention time, peak shape, integration, qualifier/quantifier behavior, internal-standard response, and the identity of preceding injections together, rather than treating peak area as an isolated number. Figure 1 shows the trend shapes and why the shape itself is the first clue.
LC-MS PK Peak Integrity Engine
Interrogate carryover, matrix effects, ion suppression, and integration integrity before your PK data goes to QC.
What do analyte and internal-standard trends tell you?
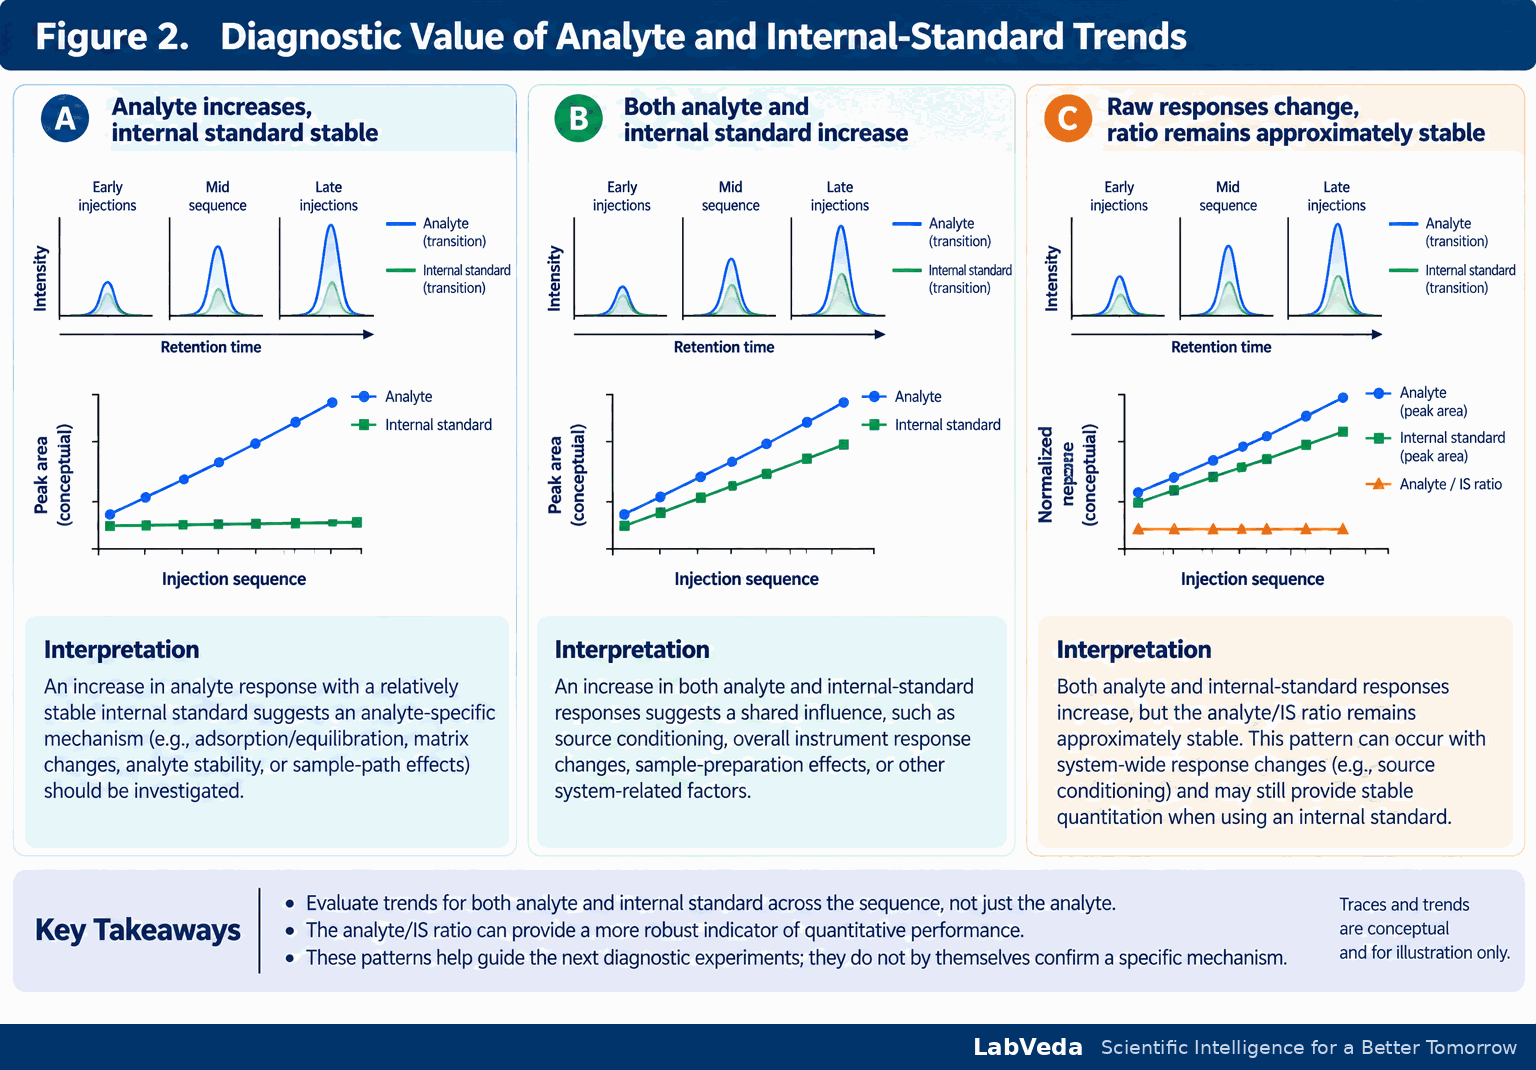
Start with the relationship between the analyte and the internal standard. If analyte response rises while the internal standard stays relatively stable, prioritize analyte-specific or sample-specific mechanisms. If both rise in parallel, investigate shared influences — system conditioning, ion-source response, LC or mobile-phase changes, or preparation effects. If the raw responses move but the analyte/internal-standard ratio stays stable, the quantitative impact may be smaller, but the underlying system change still deserves investigation. Figure 2 pairs each trend with the interpretation it supports.
What are the main mechanisms behind rising peak areas?
Several distinct mechanisms produce a similar upward trend. Table 1 summarizes the signatures that help tell them apart before any experiment is run.
| Mechanism | Typical trend signature | Neat vs matrix | Analyte / IS pattern |
|---|---|---|---|
| System or source conditioning | Rise that approaches a plateau, often from the first injections | Occurs in neat standards too | Analyte and IS often rise together; ratio relatively stable |
| Nonspecific adsorption / equilibration | Low early recovery that rises as surfaces condition | Either; depends on the surface involved | Often analyte-specific |
| Carryover / memory | Response depends on the preceding injection | Either | Selective; a following blank shows analyte |
| Matrix-dependent ionization | Rise appears in extracted samples | Matrix samples only | Analyte and IS may differ |
| Sample prep, stability or vial effects | Depends on time in the autosampler or on preparation | Usually prepared / matrix samples | Often analyte-specific |
| Instrument response / acquisition | Broad response change, tracked by QC trend | Both | System-wide; both move |
System or source conditioning
Early injections can differ from later ones while surfaces, source conditions, or other system states approach a reproducible operating condition. A rise that approaches a plateau and also occurs in neat standards is more consistent with a system-wide effect than with a matrix-only mechanism. “Conditioning” stays a hypothesis until a sequence experiment supports it.
Nonspecific adsorption and equilibration
Adsorption can occur in containers, vials, tubing, injectors, columns and other wetted surfaces. Adsorption-prone analytes may show low initial recovery that changes as surfaces become conditioned, and the location matters because mitigation differs for sample-preparation losses versus LC-system losses. Nonspecific adsorption is well documented for peptides, proteins and other adsorption-prone analytes,1 and nonspecific adsorption to LC hardware is a recognized, reviewable failure mode in its own right.2
Carryover or memory from preceding injections
Carryover is sequence-dependent: material from a preceding injection contributes to a later response, so a high sample followed by one or more blanks is a high-value diagnostic experiment. Carryover can originate from the injector, column or other flow-path components, and its quantitative impact depends on the relationship between consecutive injections.34 The mechanics of finding and washing it out are covered in the autosampler carryover guide.
Matrix-dependent ionization
If the upward trend appears in extracted matrix samples but not neat standards, investigate changing matrix effects, sample-preparation behavior, co-elution, or accumulation of matrix-derived material. In regulated bioanalysis, matrix effect is a distinct validation element and should not be conflated with carryover, stability or recovery.5 The mechanisms and the neat/post-extraction/pre-extraction experiments are treated in depth in the guide to ion suppression and matrix effects.
Sample preparation, stability or vial effects
A sequence trend can be created outside the instrument. Time in the autosampler, evaporation, adsorption to vial surfaces, incomplete mixing, precipitation, extraction variability, or analyte conversion can change the amount presented to the LC-MS/MS system6 — reinjecting the same prepared sample at different sequence positions helps separate time and order effects from between-sample preparation variability.
Instrument-response or acquisition effects
Changes in source cleanliness, tuning, detector response, gas delivery, LC composition, acquisition or integration can all alter measured response. Review system and QC trends, pressure and retention behavior, source status, calibration, the acquisition method and integration before attributing the trend to chemistry; a steady loss of response across a batch is the mirror-image problem covered in the LC-MS sensitivity-loss checklist.
How do you run experiments that discriminate the cause?
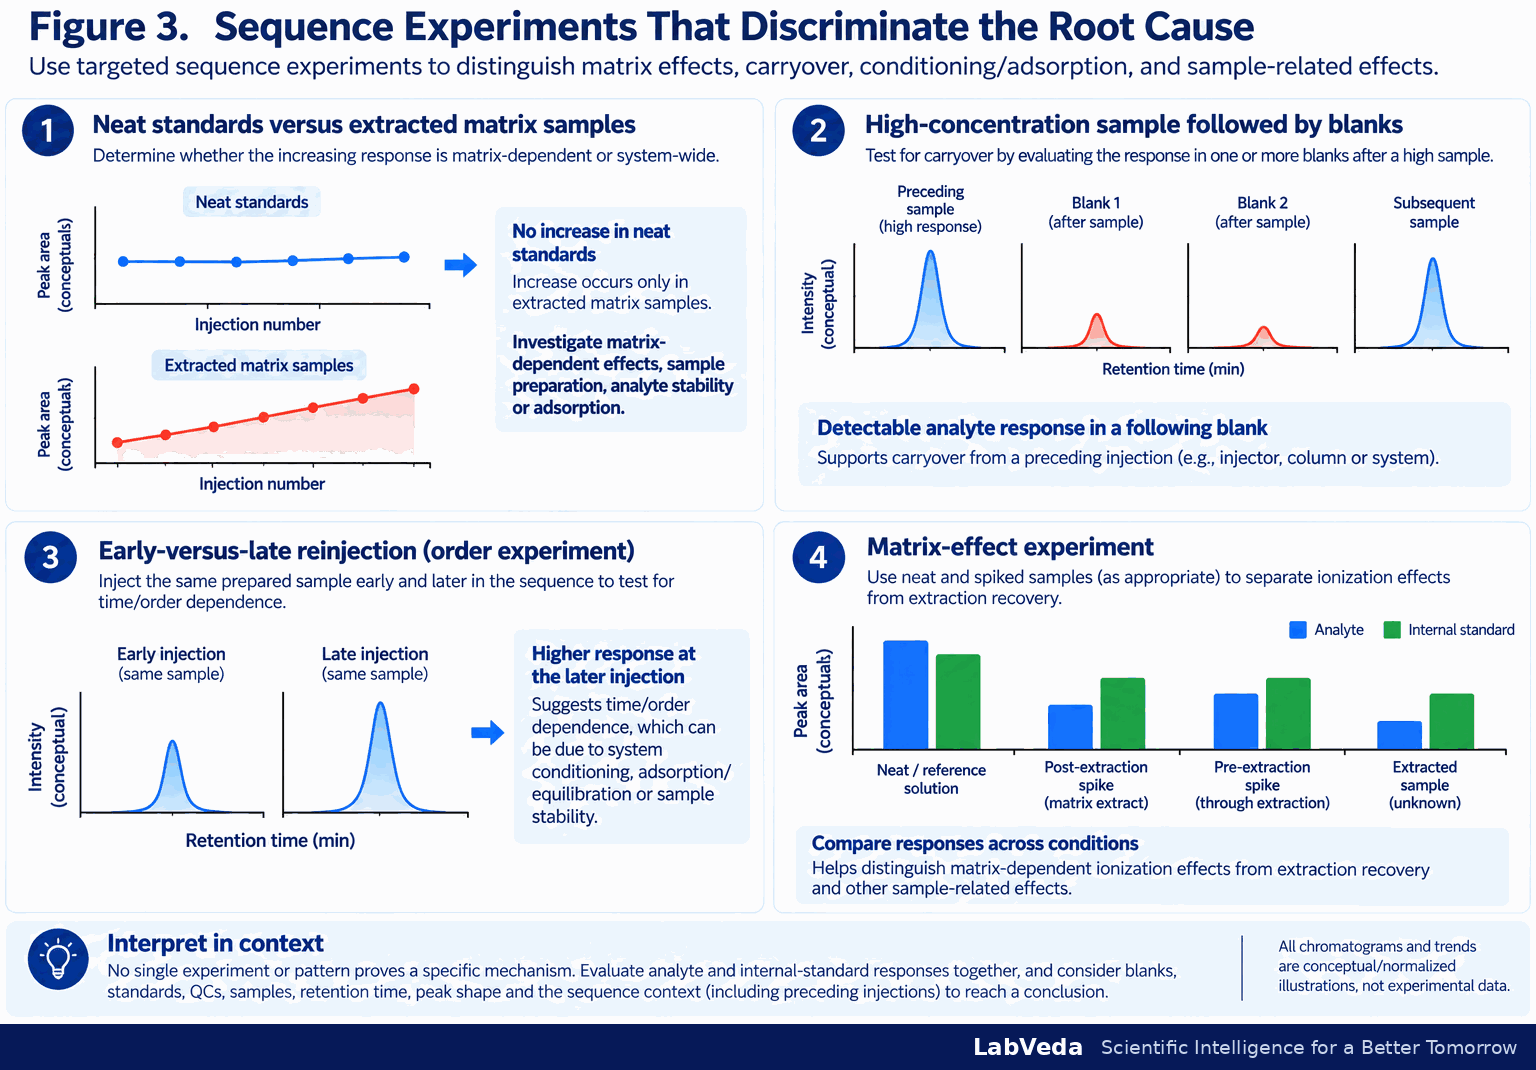
A useful diagnostic experiment changes one informative dimension while preserving the rest of the method. The objective is not to make the symptom disappear by trial and error; it is to produce an observation that meaningfully changes the probability of competing explanations. Figure 3 shows four such experiments.
Which experiment answers which question?
Table 2 pairs each diagnostic question with the experiment that answers it and the observation that supports each conclusion. A positive observation narrows the hypothesis set; it is not confirmation on its own.
| Question | Experiment | Positive observation | What it supports |
|---|---|---|---|
| Matrix-dependent? | Compare neat standards with extracted matrix samples. | Only matrix samples rise. | Matrix effect, preparation, stability, adsorption or matrix accumulation. |
| Preceding-injection dependent? | High sample → blank(s) → lower sample. | Blank contains analyte response and/or the following sample is elevated. | Carryover / memory; localize injector, column and flow path. |
| Time / order dependent? | Inject the same prepared sample early and late. | Same preparation gives a different response by position. | Conditioning / equilibration, time-dependent sample behavior or adsorption. |
| Analyte and IS coupled? | Trend analyte, IS and the analyte/IS ratio. | Both rise similarly; ratio relatively stable. | Shared system / source / preparation influence. |
| Analyte-specific? | Compare analyte and IS across standards, QCs and samples. | Analyte rises while IS is stable. | Analyte-specific adsorption, stability, matrix interaction or selective carryover. |
| Chromatography-linked? | Trend RT, peak shape, pressure and response. | Response change coincides with an LC change. | LC composition, equilibration, column or system state. |
A practical diagnostic workflow
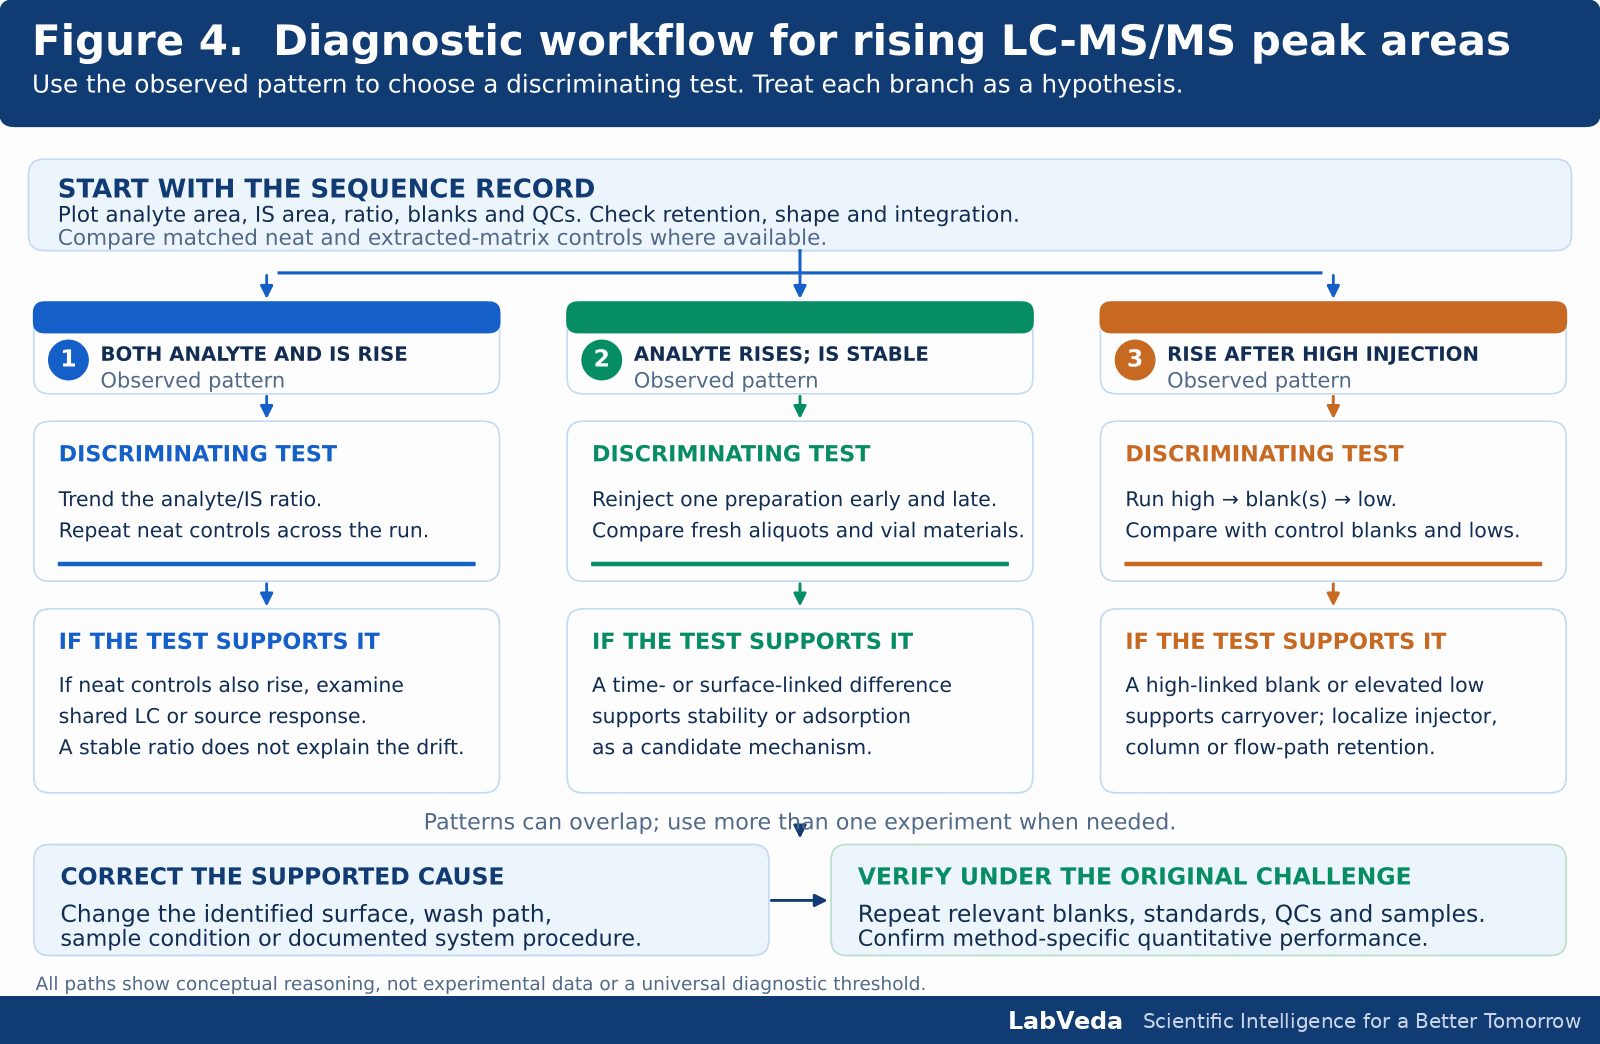
Figure 4 sequences these checks as a hypothesis-narrowing path: start from the sequence record, read the observed pattern, choose the discriminating test, and only then act. A branch says what to investigate next; it is not proof that the listed mechanism is present, and multiple mechanisms can coexist — adsorption, carryover and matrix effects can interact.
How do you correct the supported cause?
Corrective action follows the evidence. If carryover is demonstrated, localize the retained analyte and evaluate wash design, injection-path cleaning, gradient and column behavior, and sequence risk. If nonspecific adsorption is supported, identify the surface or preparation step responsible before changing materials, solvents, additives or hardware. If the effect is matrix-dependent, investigate separation, sample cleanup, dilution where scientifically appropriate, internal-standard suitability and ionization conditions. If system-wide conditioning is demonstrated, define and verify a reproducible conditioning procedure rather than relying on an undocumented number of sacrificial injections.
How do you verify the problem is actually fixed?
A successful intervention should remove or control the sequence dependence under conditions that challenge the original failure mode. Repeat the relevant standards, matrix samples, blanks, QCs and the sequence-order challenge, and confirm quantitative performance, retention behavior, internal-standard response and the method’s acceptance criteria. Do not declare success solely because one chromatogram looks better.
Where does ICH M10 fit?
ICH M10 applies to bioanalytical method validation and study-sample analysis within its stated scope; it is not a universal LC-MS troubleshooting standard. For chromatographic bioanalytical methods it treats matrix effect, calibration response and range, accuracy, precision, carryover, dilution integrity, stability and reinjection reproducibility as distinct validation elements.5 That separation is diagnostically useful: an increasing sequence trend should not be labeled “matrix effect” or “carryover” without evidence.
Frequently asked questions
Why do my LC-MS/MS peak areas keep increasing with every injection?
Possible mechanisms include system or source conditioning, analyte adsorption followed by surface equilibration, matrix-dependent effects, sample or vial changes, carryover or memory, and broader response drift. The trend shape and the analyte/internal-standard behavior determine which experiment should come next.
Does increasing peak area mean the source is becoming conditioned?
Not necessarily. A rise in neat standards that approaches a plateau can support that hypothesis, but adsorption, LC-system equilibration and other response changes can look similar.
Can carryover cause a sequence trend?
Yes, but demonstrate dependence on the preceding injections. Run blanks after challenging injections and evaluate the relationship to the preceding response rather than relying on trend shape alone.
What if the analyte and internal standard both increase?
A shared influence becomes more plausible, especially if the analyte/internal-standard ratio stays relatively stable. Investigate system or source conditioning, LC and mobile-phase changes, and preparation effects.
Should I just add more conditioning injections?
Only after evidence shows conditioning is the relevant mechanism and a reproducible conditioning procedure can be defined. Extra injections can mask the symptom without controlling the cause.
The takeaway
A rising LC-MS/MS peak area across a sequence is a symptom shared by several mechanisms — system conditioning, adsorption and equilibration, carryover, matrix-dependent ionization, sample or vial effects, and instrument-response drift — so the response itself is never the diagnosis. Read the trend shape, compare the analyte with the internal standard, test whether the rise appears in neat standards or only in matrix, and run the one experiment that separates the leading hypotheses before changing anything. Correct only the mechanism the evidence supports, then verify that the sequence dependence is gone under a challenge that would have reproduced the original failure.
References
- “Strategies to reduce aspecific adsorption of peptides and proteins in liquid chromatography-mass spectrometry based bioanalyses: an overview,” Bioanalysis (2014). PMID 25022477.
- G. J. Guimaraes, M. G. Bartlett, “Managing nonspecific adsorption to liquid chromatography hardware: a review,” Anal Chim Acta 1250, 340994 (2023). doi:10.1016/j.aca.2023.340994.
- W. Zeng et al., “A new approach for evaluating carryover and its influence on quantitation in HPLC and tandem mass spectrometry assay,” Rapid Commun Mass Spectrom (2006). PMID 16444681.
- A. Clouser-Roche, K. Johnson, D. Fast, D. Tang, “Beyond pass/fail: evaluating the effect of carryover in bioanalytical LC/MS/MS methods,” J Pharm Biomed Anal 47(1), 146–155 (2008). PMID 18242037.
- ICH, M10 Bioanalytical Method Validation and Study Sample Analysis, final guideline (2022). FDA final guidance: fda.gov.
- W. Li, S. Luo, H. T. Smith, F. L. S. Tse, “Prevention and recovery of lost analyte due to container surface adsorption in LC-MS/MS,” J Chromatogr B 878(5–6), 583–589 (2010). PMID 20097141.
Reviewed against primary sources. The mechanisms, experiments and interpretations on this page are checked against the primary literature and ICH M10 cited above. The figures are conceptual illustrations, not experimental data, and no universal threshold for an acceptable sequence trend is implied. For validated or compendial methods, the applicable procedure and regulatory framework take precedence over the general guidance given here. Evidence review: September 2026.